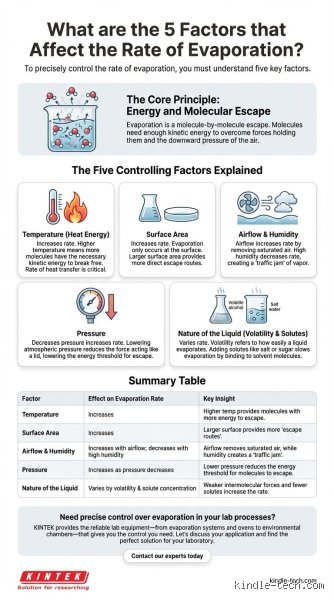
Deciphering the thermodynamics of Phase Change: On what factors does the rate of Evaporation depend?
At the microscopic level, liquid molecules are bound together by intermolecular forces such as hydrogen bonding, dipole-dipole interactions, and London dispersion forces. These forces create a cohesive net holding the liquid phase together. However, thermal energy causes these molecules to constantly vibrate, collide, and exchange energy. When an individual molecule acquires sufficient kinetic energy to overcome these cohesive intermolecular bonds near the liquid-gas boundary, it breaks free from the liquid matrix and enters the gas phase as vapor.
The overall speed at which this transition happens depends on a set of core environmental and intrinsic properties. This first installment of our expert guide explores the primary variables dictating evaporation kinetics, beginning with thermal energy and surface exposure.
Temperature and kinetic energy distribution
The Maxwell-Boltzmann distribution and Vapor Pressure
Temperature serves as the most potent driver of evaporation rates. Macroscopically, temperature is a measure of the average translational kinetic energy possessed by the particles within a substance. Microscopically, however, the molecules in a liquid do not all share the exact same energy. Instead, their energy follows a statistical distribution known as the Maxwell-Boltzmann distribution.
Within this distribution, a small fraction of molecules possesses significantly higher kinetic energy than the average, while another fraction possesses lower energy. Only those molecules positioned at the high-energy tail of the curve possess enough velocity to sever the intermolecular attractions binding them to their neighbors.
Direct Proportionality: As the system temperature increases, the entire distribution curve shifts toward higher energy states. Consequently, a substantially larger population of molecules exceeds the escape threshold per unit of time.
Vapor Pressure Elevation: Higher temperatures increase the vapor pressure of the liquid—the pressure exerted by its vapor in thermodynamic equilibrium with its condensed phases. As vapor pressure rises, molecules escape into the surrounding atmosphere with greater frequency.
Adding thermal energy to a system also accelerates molecular motion within the boundary layer right above the liquid surface, reducing the density of the gas phase immediately adjacent to the interface and facilitating faster outward diffusion.
Surface area and Interfacial exposure
Geometric constraints and Macro-Scale dynamics
While thermal energy dictates the energy of individual particles, surface area determines how many particles have physical access to the escape boundary. Evaporation is strictly an interfacial phenomenon; it occurs exclusively at the boundary where the liquid meets the gas phase. Molecules residing deep within the bulk liquid cannot escape directly into the atmosphere because they are surrounded on all sides by neighboring molecules exerting inward cohesive forces.
Interfacial Access: Expanding the surface area of a liquid increases the spatial footprint available for molecular departure. For instance, spreading fifty milliliters of water across a wide shallow tray causes it to evaporate exponentially faster than keeping the same volume concentrated inside a narrow test tube.
Collision Probability: A larger surface area maximizes the number of boundary-facing molecules exposed to ambient thermal radiation and air currents, raising the statistical probability of successful molecular escape events per second.
Industrial applications regularly exploit this principle. Cooling towers, desalination plants, and chemical distillation setups utilize spray nozzles, misting arrays, or packed beds to atomize liquids into microscopic droplets or spread them across thin films. This mechanical intervention maximizes surface area-to-volume ratios, optimizing mass transfer and accelerating evaporation rates by orders of magnitude compared to static pools.
Comparing Evaporation and Boiling: Alternative Phase Change mechanisms
Surface vs. Bulk vaporization characteristics
To fully comprehend the parameters governing evaporation, it helps to compare it directly against boiling, an alternative phase change mechanism that converts liquid to gas. While both processes involve liquid-to-gas phase transitions, their underlying thermodynamic constraints differ significantly.
| Feature | Evaporation | Boiling |
|---|---|---|
| Location of Phase Change | Exclusively at the liquid-gas surface | Throughout the bulk liquid (including interior) |
| Temperature Requirement | Occurs at any temperature below the boiling point | Occurs strictly at or above the boiling point |
| Vapor Pressure Relationship | Vapor pressure is less than ambient atmospheric pressure | Vapor pressure equals or exceeds ambient atmospheric pressure |
| Bubble Formation | Absent; molecules escape individually from the surface | Present; vapor bubbles form within the bulk liquid and rise |
Evaporation happens quietly and incrementally across a wide range of temperatures, driven entirely by the ambient humidity gradient and surface thermal energy. Boiling, conversely, is a violent and rapid phase change that requires the vapor pressure of the liquid to match or exceed the external pressure exerted upon it by the atmosphere. At the boiling point, thermal energy is high enough to generate vapor bubbles anywhere within the liquid volume, rather than just at the surface boundary.
Understanding this distinction clarifies why damp clothing dries outdoors on a chilly winter day—evaporation proceeds efficiently as long as the surrounding air is not saturated, even though the temperature remains far below the boiling point of water.
Molecular kinetics and boundary layer dynamics
Moving beyond the macroscopic observations of temperature and surface area, the true engine of evaporation lies in statistical mechanics and fluid dynamics at the microscopic interface. At any given moment, the molecules within a liquid are not possessing a uniform energy state; instead, they exist in a continuous distribution governed by the Maxwell-Boltzmann distribution.
Within this statistical spread, a small fraction of molecules possess kinetic energies significantly higher than the system's average. When these high-energy molecules migrate toward the liquid-gas boundary, they must overcome the cohesive intermolecular forces holding the liquid together, such as hydrogen bonding, dipole-dipole interactions, or London dispersion forces. This energy barrier is directly proportional to the substance's enthalpy of vaporization. Only those molecules whose outward vector and kinetic energy surpass this threshold can break free from the liquid surface, transitioning into the vapor phase.
However, escape is only the first step. Once a molecule enters the gas phase immediately adjacent to the liquid surface, it contributes to the local vapor pressure. If the surrounding air is stagnant, these newly liberated molecules accumulate, forming a localized vapor boundary layer. This micro-environment establishes a concentration gradient that drives back-diffusion; molecules collide with air particles and have a statistical probability of re-entering the liquid via condensation.
To sustain a high evaporation rate, mass transfer must overcome this boundary layer resistance. This is where fluid dynamics and ambient airflow become critical. Convective mass transfer sweeps away saturated vapor parcels, maintaining a steep concentration gradient between the liquid surface and the bulk atmosphere. Engineers and physicists model this using dimensionless numbers such as the Sherwood number, Reynolds number, and Schmidt number, which quantify the interplay between forced/natural convection, molecular diffusion, and kinematic viscosity. When wind or forced ventilation disrupts the boundary layer, the resistance to mass transfer plummets, causing the evaporation rate to scale dramatically.
Little-Known aspect: Curvature, Pressure, and Knudsen Evaporation
While standard textbooks emphasize temperature, humidity, and wind, advanced thermodynamic frameworks reveal subtle variables that drastically alter evaporation dynamics under specific conditions:
Surface Curvature and the Kelvin Effect: Flat surfaces exhibit a standard equilibrium vapor pressure. However, when liquids form micro-droplets, curved menisci, or exist within sub-micron porous media, the local vapor pressure changes significantly. The Kelvin equation dictates that convex surfaces (like tiny droplets) have a higher equilibrium vapor pressure than flat surfaces, making them evaporate much faster than expected. Conversely, concave menisci in capillary tubes lower vapor pressure, suppressing evaporation.
The Role of Total Atmospheric Pressure: A common misconception is that atmospheric pressure only matters at the boiling point. In reality, total pressure influences the diffusion coefficient (DABD_{AB}DAB) of vapor molecules moving through air. According to the Chapman-Enskog theory derived from kinetic gas theory, binary diffusion coefficients are inversely proportional to total pressure. In a partial vacuum, molecules experience fewer collisional interruptions, allowing vapor to diffuse away from the interface at accelerated rates even if the thermal energy remains constant.
Knudsen Regimes in Microfluidics: When the characteristic dimensions of the evaporation system approach the mean free path of the evaporating molecules, such as in nanotechnology or nanoporous membranes, the continuous fluid dynamics model breaks down entirely. This is known as the Knudsen regime. Here, molecules collide with the pore walls more frequently than with each other, rendering traditional mass transfer equations obsolete and introducing distinct evaporation kinetics driven strictly by molecular-wall collision mechanics.
Final Synthesis
Understanding evaporation requires recognizing that it is not merely a surface phenomenon, but a complex interplay between statistical molecular energy distribution, interfacial thermodynamics, and boundary-layer mass transport. While environmental drivers like temperature, humidity, and airflow dictate macroscopic rates in everyday scenarios, microscopic factors such as molecular geometry, local pressure fields, and surface curvature ultimately govern phase change at the fundamental level. Therefore, controlling or predicting evaporation in industrial, meteorological, or biological systems demands a holistic approach that bridges microscopic kinetics with macroscopic fluid dynamics.