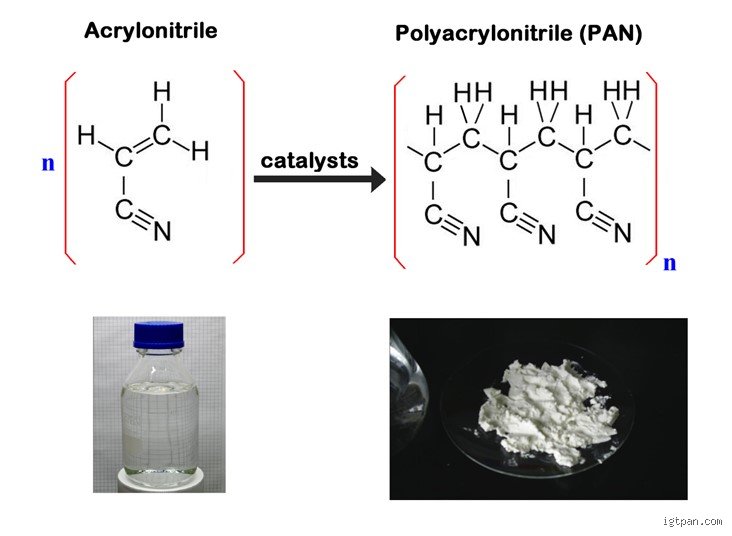
Understanding the structural integrity, mechanical performance, and thermal conversion behavior of PAN requires a rigorous examination of the chemical reaction responsible for its formation. The polymerization of acrylonitrile is a classic yet uniquely nuanced example of chain-growth addition polymerization, heavily characterized by precipitation phenomena, strong dipole-dipole interactions, and specific stereochemical outcomes.
1. The Monomer foundation: Chemical architecture of acrylonitrile
The foundational building block of polyacrylonitrile is acrylonitrile (CH2=CH−CNCH_2=CH-CNCH2=CH−CN), an organic compound consisting of a vinyl group bonded directly to a strongly electronegative cyano group (−C≡N- ext{C}\equiv ext{N}−C≡N). Structurally, acrylonitrile is an asymmetrical molecule featuring a carbon-carbon double bond (C=C ext{C}= ext{C}C=C) adjacent to an electron-withdrawing nitrile functional group.
This specific molecular arrangement dictates the electronic distribution across the π\piπ-bond system. The strong electron-withdrawing inductive and resonance effects of the cyano group polarize the adjacent vinyl bond, lowering the electron density on the β\betaβ-carbon and rendering the double bond exceptionally susceptible to attack by free radicals. The compact size and high polarity of the nitrile group heavily influence the intermolecular forces of the resulting polymer, imparting rigidity, high melting resistance, and remarkable chemical inertness to the macroscopic polymer matrix.
2. Primary Reaction pathway: Free-Radical addition Polymerization
The industrial and laboratory synthesis of polyacrylonitrile proceeds almost exclusively via chain-growth addition polymerization (specifically, free-radical polymerization). While ionic mechanisms (such as anionic polymerization) can theoretically be initiated under specialized cryogenic or strictly anhydrous conditions, cationic polymerization is virtually non-existent due to the electron-withdrawing nature of the cyano group, which destabilizes any developing positive charge on the carbon backbone.
In standard industrial practice, free-radical polymerization is favored because it accommodates diverse manufacturing setups, allows precise control over molecular weights through chain transfer agents, and scales efficiently. The overall reaction converts thousands of individual monomer units into a high-molecular-weight macromolecule characterized by a saturated carbon backbone adorned with pendant nitrile groups:
n CH2=CH(CN)→Initiator/Energy −[CH2−CH(CN)]n−n \, ext{CH}_2= ext{CH}( ext{CN}) \xrightarrow{ ext{Initiator/Energy}} \, -[ ext{CH}_2- ext{CH}( ext{CN})]_n-nCH2=CH(CN)Initiator/Energy−[CH2−CH(CN)]n−
Because the propagation step involves the sequential opening of π\piπ-bonds to form stable σ\sigmaσ-bonds, the reaction is strongly exothermic. Managing this thermal output is a critical engineering challenge in commercial reactors to prevent localized overheating and unwanted secondary reactions.
3. Step-by-Step Reaction mechanism
Like all classical free-radical polymerizations, the formation of polyacrylonitrile proceeds through three fundamental kinetic stages: Initiation, Propagation, and Termination.
A. Initiation
The initiation phase requires the generation of reactive free radicals capable of attacking the carbon-carbon double bond of the acrylonitrile monomer. This is typically achieved using thermal or redox initiators. Common thermal initiators include azo compounds like azobisisobutyronitrile (AIBN) or peroxides such as benzoyl peroxide. In aqueous or suspension systems, redox initiating systems, such as persulfate-bisulfite combinations, are frequently employed.
Homolytic Cleavage: The initiator molecule undergoes thermal or chemical homolytic fission, breaking a covalent bond to yield two active radical species (R∙R^\bulletR∙).
Monadic Attack: One of these newly formed radicals attacks the π\piπ-electron cloud of an acrylonitrile monomer molecule. The radical adds to the less substituted carbon (CH2 ext{CH}_2CH2) of the vinyl group, shifting the unpaired electron to the more substituted adjacent carbon atom. This generates an active chain-initiating radical species:
R∙+CH2=CH(CN)⟶R−CH2−C˙H(CN)R^\bullet + ext{CH}_2= ext{CH}( ext{CN}) \longrightarrow R- ext{CH}_2-\dot{ ext{C}} ext{H}( ext{CN})R∙+CH2=CH(CN)⟶R−CH2−C˙H(CN)
B. Propagation
Once the initial radical adduct is formed, the propagation stage drives the rapid, continuous growth of the polymer chain. The newly generated carbon-centered radical attacks the double bond of another incoming acrylonitrile monomer. The π\piπ-bond opens, adding the monomer unit to the growing chain while regenerating the active radical center at the terminal end:
R−(CH2−CH(CN))n−CH2−C˙H(CN)+CH2=CH(CN)⟶R−(CH2−CH(CN))n+1−CH2−C˙H(CN)R-( ext{CH}_2- ext{CH}( ext{CN}))_n- ext{CH}_2-\dot{ ext{C}} ext{H}( ext{CN}) + ext{CH}_2= ext{CH}( ext{CN}) \longrightarrow R-( ext{CH}_2- ext{CH}( ext{CN}))_{n+1}- ext{CH}_2-\dot{ ext{C}} ext{H}( ext{CN})R−(CH2−CH(CN))n−CH2−C˙H(CN)+CH2=CH(CN)⟶R−(CH2−CH(CN))n+1−CH2−C˙H(CN)
This step occurs at an extraordinarily rapid rate. Because the cyano groups stabilize the free radical intermediate through resonance structures involving the nitrogen lone pair, the propagation rate constant remains robust across various solvent environments.
C. Termination
Polymerization cannot continue indefinitely; it must conclude via chain termination, which halts the growth of the macromolecule. Termination in PAN synthesis typically proceeds via one of two primary pathways:
Combination (Coupling): Two active growing polymer chains encounter one another, and their terminal radical electrons pair up to form a single covalent carbon-carbon bond, yielding a polymer chain with double the kinetic chain length and specific terminal groups.
Disproportionation: A hydrogen atom is abstracted from one growing radical chain by another radical chain. This results in the formation of two distinct dead polymer molecules: one saturated chain and one terminal chain possessing an unsaturated terminal alkene group.
4. The unique phenomenon of Precipitation Polymerization
A defining characteristic that sets the formation of polyacrylonitrile apart from standard vinyl polymers like polystyrene or polymethyl methacrylate is its physical phase behavior. Polyacrylonitrile is exceptionally insoluble in its own monomer and exhibits very limited solubility in many common organic solvents and water.
Consequently, the synthesis of PAN generally proceeds via precipitation polymerization (a form of heterogeneous polymerization).
Homogeneous Initiation: The reaction begins in a homogeneous phase where the initiator and monomer are dissolved in a suitable solvent (such as water, dimethylformamide, dimethyl sulfoxide, or aqueous mixtures).
Phase Separation: As the propagation phase proceeds and the molecular weight of the growing polymer chains exceeds a critical threshold (typically only a few repeating units), the polymer loses solubility in the reaction medium.
Micro-particle Precipitation: The polymer chains collapse and precipitate out of the solution as discrete micro-particles or aggregates.
Despite precipitating out, these polymer particles often retain trapped radicals or remain accessible to dissolved monomers, allowing polymerization to continue within the swollen polymer phase (occlusion effect). This heterogeneous nature introduces unique kinetic complexities, such as autoacceleration (the Trommsdorff effect), where termination rates drop sharply because the radical chain ends are physically restricted inside the precipitated solid matrix.
Chain-Growth Polymerization mechanics: Initiation, Propagation, and Termination
Because acrylonitrile possesses a highly electron-deficient carbon-carbon double bond, resulting from the strong electron-withdrawing inductive and resonance effects of the pendant nitrile (−C≡N- ext{C}\equiv ext{N}−C≡N) group, its polymerization proceeds predominantly via a free-radical chain-growth mechanism. While anionic polymerization is theoretically viable using strong nucleophilic initiators such as alkyllithiums or sodium amide, industrial manufacturing relies almost exclusively on free-radical pathways due to their superior tolerance for trace protic impurities and compatibility with aqueous reaction media.
The free-radical polymerization of acrylonitrile (CH2=CH−CNCH_2=CH-CNCH2=CH−CN) unfolds through three distinct kinetic stages:
Initiation: The reaction begins with the homolytic cleavage of a thermally unstable initiator molecule to generate primary free radicals (R∙R\bulletR∙). In commercial solution or suspension processes, azo compounds, most notably azobisisobutyronitrile (AIBN), or organic peroxides such as benzoyl peroxide are utilized. In aqueous redox systems, combinations of persulfates (e.g., potassium persulfate, K2S2O8K_2S_2O_8K2S2O8) and bisulfites (NaHSO3NaHSO_3NaHSO3) generate sulfate and bisulfite radicals at lower operating temperatures (typically 40°C to 60°C). Once formed, the primary radical attacks the less sterically hindered β\betaβ-carbon (methylene carbon) of the acrylonitrile monomer, breaking the π\piπ-bond and transferring the unpaired electron to the α\alphaα-carbon, which is stabilized by resonance delocalization across the adjacent nitrile triple bond:
R∙+CH2=CH(CN)⟶R−CH2−C˙H(CN)R\bullet + CH_2=CH(CN) \longrightarrow R-CH_2-\dot{C}H(CN)R∙+CH2=CH(CN)⟶R−CH2−C˙H(CN)Propagation: The newly formed monomeric radical acts as a reactive center, rapidly adding successive acrylonitrile units in a head-to-tail regioselective orientation. Because the nitrile group stabilizes the propagating radical on the α\alphaα-carbon and minimizes steric repulsion during approach, head-to-head additions are thermodynamically and kinetically disfavored. The propagation step repeats thousands of times within fractions of a second, building a high-molecular-weight macromolecular backbone:
R−[CH2−CH(CN)]n−CH2−C˙H(CN)+CH2=CH(CN)⟶R−[CH2−CH(CN)]n+1−CH2−C˙H(CN)R-[CH_2-CH(CN)]_n-CH_2-\dot{C}H(CN) + CH_2=CH(CN) \longrightarrow R-[CH_2-CH(CN)]_{n+1}-CH_2-\dot{C}H(CN)R−[CH2−CH(CN)]n−CH2−C˙H(CN)+CH2=CH(CN)⟶R−[CH2−CH(CN)]n+1−CH2−C˙H(CN)Termination and Chain Transfer: Chain growth ceases when two active macroradicals encounter one another. For polyacrylonitrile (PAN), termination occurs predominantly via combination (coupling of two growing chains to form a single inactive polymer with a head-to-head linkage at the juncture) rather than disproportionation, particularly at moderate reaction temperatures. However, a defining kinetic quirk of acrylonitrile polymerization is the high rate of chain transfer to monomer and solvent. In homogeneous solutions, solvents like dimethylformamide (DMF) or dimethyl sulfoxide (DMSO) possess abstractable hydrogen atoms; a propagating radical may abstract a hydrogen atom from a solvent molecule, terminating the original polymer chain while generating a new solvent radical capable of initiating a fresh chain.
The heterogeneous Precipitation phenomenon and autoacceleration
A critical feature distinguishing the bulk and aqueous polymerization of acrylonitrile from that of styrene or methyl methacrylate is the insolubility of the resulting polymer in its own monomer and in water. Polyacrylonitrile exhibits intense intermolecular dipole-dipole interactions between adjacent nitrile groups (which possess a high dipole moment of approximately 3.9 D3.9 ext{ D}3.9 D), resulting in a tightly packed, pseudo-crystalline morphology with a high cohesive energy density.
Consequently, as soon as oligomeric radicals reach a critical degree of polymerization, typically around 10 to 20 monomer units, they precipitate out of the continuous liquid phase, coalescing into swollen, solid polymer particles. This phase separation fundamentally alters the reaction kinetics through the Trommsdorff-Norrish effect (gel effect), manifesting in a pronounced autoacceleration of the polymerization rate:
Diffusion-Controlled Termination: Once the growing radical chains become occluded within the viscous, precipitated polymer particles, their translational mobility drops precipitously. Two bulky macroradicals can no longer diffuse freely to meet and terminate, causing the termination rate constant (ktk_tkt) to plummet by several orders of magnitude.
Unimpeded Propagation: Small, uncharged acrylonitrile monomer molecules continue to diffuse readily through the swollen polymer matrix to reach the trapped active radical sites. Thus, the propagation rate constant (kpk_pkp) remains largely unaffected until very high conversions are achieved.
Kinetic Consequence: Because the overall rate of polymerization (RpR_pRp) is proportional to kp/ktk_p / \sqrt{k_t}kp/kt, the sharp decline in ktk_tkt triggers a surge in radical concentration, driving an exponential increase in reaction velocity and exothermic heat release (ΔH≈−76.5 kJ/mol\Delta H \approx -76.5 ext{ kJ/mol}ΔH≈−76.5 kJ/mol). Without rigorous thermal management, this autoacceleration can lead to thermal runaway.
Industrial Polymerization methodologies
To manage the high exothermicity and insolubility of PAN, chemical engineers employ specific reactor configurations tailored to the intended end-use of the polymer, primarily textile fibers or carbon fiber precursors.
| Process Type | Reaction Medium | Initiator System | Key Operational Characteristics | Primary Application |
|---|---|---|---|---|
| Aqueous Suspension (Slurry) | Deionized water | Redox (Persulfate / Bisulfite) | Monomer is partially soluble (~7% at 50°C); polymer precipitates as a fine filterable powder. Easy heat removal. | General-purpose acrylic fibers; dry/wet spinning after dissolution. |
| Solution Polymerization | Polar aprotic solvents (DMF, DMSO, DMAc) or concentrated inorganic salts (NaSCN, ZnCl2) | Thermal free-radical (AIBN) | Homogeneous single-phase system; eliminates precipitation and enables direct spinning without polymer isolation. | High-performance textiles and carbon fiber precursors. |
| Emulsion Polymerization | Water with anionic surfactants | Water-soluble persulfates | Produces stable latex dispersions; high molecular weights achieved rapidly. | Specialty coatings, binders, and impact-modified copolymers. |
Copolymerization: Overcoming homopolymer limitations
Pure polyacrylonitrile homopolymer is virtually unprocessable by conventional melt-extrusion techniques. Because the strong dipolar forces between nitrile groups hold the polymer chains together with immense thermal stability, PAN undergoes cyclization and thermal degradation at temperatures around 250∘C250^\circ ext{C}250∘C to 300∘C300^\circ ext{C}300∘C, well below its theoretical crystalline melting point of approximately 317∘C317^\circ ext{C}317∘C. Even when dissolved in polar aprotic solvents for wet or dry spinning, pure PAN yields fibers that are overly rigid, brittle, and impervious to standard textile dyes.
To overcome these limitations, commercial "polyacrylonitrile" is almost universally synthesized as a copolymer containing between 85% and 99% acrylonitrile by weight, modified with 1% to 15% of carefully selected comonomers:
Neutral Plasticizing Comonomers: Monomers such as methyl acrylate or vinyl acetate are incorporated at 5% to 10% levels. These bulky, non-polar or moderately polar side groups act as internal spacers, disrupting the regular packing of the nitrile dipoles. This lowers the glass transition temperature (TgT_gTg), increases chain segment mobility, and improves solubility in spinning solvents.
Ionic Dye-Site Comonomers: For textile applications, small quantities (0.5% to 2%) of acidic or basic comonomers, such as sodium styrene sulfonate, itaconic acid, or 2-methyl-5-vinylpyridine, are copolymerized into the backbone. These functional groups introduce permanent anionic or cationic sites capable of binding basic (cationic) or acid dyes, respectively.
Stabilization Accelerators (Carbon Fiber Precursors): When PAN is destined for conversion into high-modulus carbon fiber, neutral comonomers like vinyl acetate are excluded because they leave voids during pyrolysis. Instead, 1% to 3% of itaconic acid or methacrylic acid is introduced. During the subsequent thermal stabilization stage (200∘C200^\circ ext{C}200∘C–300∘C300^\circ ext{C}300∘C in air), the carboxylic acid groups initiate an ionic cyclization of the adjacent nitrile groups at lower temperatures, broadening the exothermic peak and preventing fiber fusion or core-shell structural defects.
Post-Polymerization chemistry: Thermal cyclization and carbonization
The chemical reactivity of the polyacrylonitrile macromolecule does not end upon completion of the polymerization reaction; rather, its pendant nitrile groups serve as the chemical foundation for manufacturing high-strength carbon fibers. When linear PAN fibers are subjected to controlled oxidation in air between 200∘C200^\circ ext{C}200∘C and 300∘C300^\circ ext{C}300∘C, the nitrile triple bonds undergo an intramolecular nucleophilic addition, converting the linear carbon-carbon backbone into a thermally stable, conjugated ladder polymer composed of fused six-membered naphthyridine rings.
Following this oxidative stabilization, the ladder polymer is heated to temperatures exceeding 1,000∘C1,000^\circ ext{C}1,000∘C to 1,500∘C1,500^\circ ext{C}1,500∘C in an inert nitrogen atmosphere (carbonization). During this phase, non-carbon heteroatoms, specifically hydrogen, nitrogen, and oxygen, are expelled as volatile gases (HCNHCNHCN, NH3NH_3NH3, H2OH_2OH2O, CO2CO_2CO2, and N2N_2N2), while the aromatic ladder structures fuse laterally into turbostratic carbon sheets aligned along the fiber axis. Thus, the precise control of molecular weight, comonomer distribution, and chain linearity achieved during the initial free-radical addition polymerization directly dictates the tensile strength and modulus of the final carbon material.