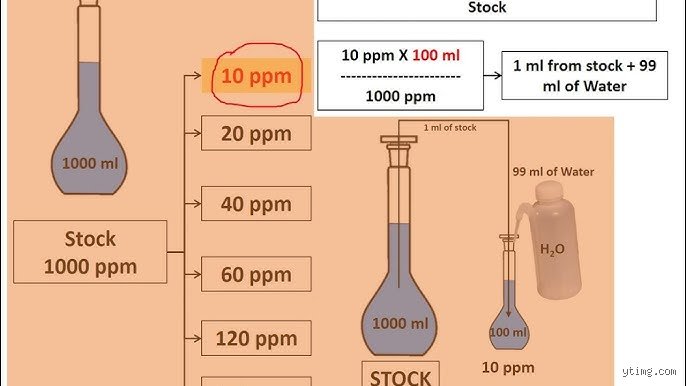
The hidden reality of parts per million calculations
A single part per million represents one milligram of a substance dissolved in a full liter of liquid. When you are handed a bottle labeled 1000 ppm, you are dealing with a relatively concentrated stock solution, often used as the baseline for calibration curves in analytical chemistry. But what does that actually mean when you step up to the workbench?
Deconstructing the volumetric ratio
Let's look at the math through a different lens because standard textbooks love to overcomplicate things. A concentration of 1000 ppm is equivalent to 1000 milligrams per liter, which can also be written as 1 gram per liter or 0.1% by weight. Dropping that down to 10 ppm means you are targeting a mere 10 milligrams per liter. The dilution factor is staring right at you. You are dividing the initial concentration by the final concentration, which gives you a factor of exactly 100. Yet the problem is that rookie technicians frequently mix up the ratio of solute to solvent, mistakenly adding 1 part stock to 100 parts solvent, which accidentally creates a 1:101 dilution and skews the final concentration to 9.9 ppm. That matters a lot if you are calibrating a high-end mass spectrometer where a 1% error margin throws off a whole week of data.
Why mass and volume can feud in high concentrations
In an ideal world, we assume that 1 milliliter of water weighs exactly 1 gram, a clean relationship that allows us to swap volumetric measurements for mass without a second thought. But room temperature fluctuates. If you are working in a sticky, non-air-conditioned facility in Mumbai during July, the density of your water drops, meaning your volumetric glassware is no longer telling the absolute truth. I have seen laboratories lose thousands of dollars in ruined agricultural assays simply because they ignored ambient temperature during a 1000 ppm to 10 ppm reduction. Most experts agree that volumetric measurements are fine for routine work, but it's unclear why more labs don't just switch to gravimetric prep—weighing everything on an analytical balance—to completely bypass the thermal expansion of liquids.
The Step-by-Step mechanics of a 1:100 serial dilution
Achieving a pristine 10 ppm solution requires more than just steady hands; it demands a strict adherence to volumetric protocol. You cannot just grab a plastic beaker and guess your way to analytical precision.
The equipment matrix for High-Accuracy pipetting
Before you even open your stock bottle, your workbench needs to look like a high-end metrology lab. You will need a Class A volumetric pipette, a calibrated 100 mL volumetric flask, and high-purity deionized water (Type 1, 18.2 megohm-cm resistance). Why choose Class A over cheaper options? Because the manufacturing tolerances are twice as strict, meaning a 1 mL Class A pipette has an error margin of just plus or minus 0.007 mL. If you use a battered, uncalibrated mechanical pipette from the 1990s, you might as well be guessing the volume by eye.
Executing the transfer without contamination
First, rinse your 1 mL pipette with a tiny amount of the 1000 ppm stock solution and discard the rinse to ensure no residual water dilutes your aliquot. Next, draw the stock solution up past the graduation mark on your pipette, then slowly drop the liquid until the bottom of the meniscus rests perfectly on the line. Transfer this 1 mL slice into your 100 mL volumetric flask, ensuring the tip touches the glass wall to draw out the last hanging drop. Now, add your diluent, whether it is pure water, 2% nitric acid for trace metal analysis, or HPLC-grade methanol, until the flask is roughly 80% full. Swirl it gently. Do not shake it violently yet, because swirling ensures the solutes mix without creating air bubbles that hide the true meniscus line. Finally, use a disposable Pasteur pipette to add the last few drops of solvent slowly until the meniscus kisses the 100 mL graduation ring perfectly. Invert the stoppered flask at least ten times to guarantee absolute homogeneity throughout the mixture.
Advanced mathematics behind the C1V1 equation
We cannot discuss how to dilute 1000 ppm to 10 ppm without paying homage to the classic conservation of mass formula. It is the bedrock of chemical preparation, even if it gives undergraduate students flashbacks to freshman chemistry exams.
Breaking down the standard dilution formula
The equation is elegant in its simplicity: $$C_1V_1 = C_2V_2$$. Here, $C_1$ represents your initial concentration ($1000 ext{ ppm}$), $V_1$ is the volume of that stock you need to find, $C_2$ is your target concentration ($10 ext{ ppm}$), and $V_2$ is the total volume you want to create. If we plug our target numbers into the formula to find out how much stock we need for a 100 mL final volume, the math rearranges beautifully: $$1000 imes V_1 = 10 imes 100$$ $$V_1 = \frac{1000}{1000}$$ $$V_1 = 1 ext{ mL}$$
But what if you need a massive 5-liter batch for a pilot plant test in a chemical facility? The formula scales perfectly, showing you would need 50 mL of stock topped up to 5000 mL. That's why this specific mathematical relationship is coded into almost every automated liquid handling system found in modern pharmaceutical laboratories.
Direct versus serial dilution: Choosing your strategy
When you need to drop a concentration by two orders of magnitude, you have a critical tactical choice to make. You can either do it in one single jump or break it down into smaller, successive steps.
The One-Step direct approach and its blind spots
Direct dilution is fast. You pull 1 mL, dump it into a 100 mL flask, dilute to the mark, and you are done. But look closely at the potential for disaster here: if your 1 mL pipette is off by just 0.02 mL due to a tiny droplet clinging to the inside of the plastic tip, your final solution shifts from 10 ppm to 10.2 ppm. That is a massive 2% error introduced in a single heartbeat! Is saving three minutes of lab time worth risking a false positive during an environmental compliance audit? For routine, low-stakes work, the direct method is acceptable, but sharp analysts know that single-step jumps over a 100-fold gap push the boundaries of manual pipetting precision.
Common pitfalls and volumetric misconceptions
The meniscus illusion and volumetric flaws
Precision hangs on a curved thread. When you aim to dilute 1000 ppm to 10 ppm, a single misplaced drop wrecks the entire calculation. Parallax error ruins accuracy because looking at a pipette from an angle alters your perceived volume. You must align your eye exactly parallel to the bottom of the liquid curve. But let's be clear: a standard glass beaker is a decorative jar, not a measurement device. Relying on beaker markings for a hundred-fold reduction guarantees failure. You need a Class A volumetric flask to hit your target concentration without introducing a massive 5% margin of error.Temperature neglect altering volume
Fluids expand. If your stock solution sits under a hot laboratory lamp, its density drops. Testing at 28°C using calculations calibrated for 20°C introduces silent, frustrating discrepancies.
Because water possesses a temperature-dependent coefficient of expansion, your volumetric measurements shift. You might think you achieved a perfect dilution, except that the physical mass of the solute per unit volume has changed.
Cross-Contamination in sequential steps
Reusing the same plastic pipette tip across multiple steps introduces unwanted solute carryover. Residual liquid clinging to the inner walls of your transfer tool will corrupt the final mixture. A mere 0.05 milliliters of leftover 1000 ppm stock can spike your final container well past the desired threshold.The logarithmic Factor and expert insights
Why serial dilution beats the Single-Step method
Why do seasoned analytical chemists reject the idea of jumping straight from the concentrate to the trace level? The problem is the sheer physical limitation of micro-pipetting. To execute a direct 1:100 reduction, you would need to introduce 1 milliliter of your concentrated material into exactly 99 milliliters of solvent. [Image of serial dilution process] $$C_1 V_1 = C_2 V_2$$ $$ ext{Dilution Factor} = \frac{1000 ext{ ppm}}{10 ext{ ppm}} = 100$$ Attempting this in one massive leap magnifies mechanical delivery errors. If your pipette misses the mark by a tiny fraction, the final variance becomes amplified. Instead, the smartest route involves a two-stage serial dilution process where you create an intermediate benchmark. First, dilute your initial stock by a 1:10 ratio to achieve a stable 100 ppm environment. After that, perform a subsequent 1:10 transfer using that fresh intermediate mixture. This geometric progression keeps the volumetric demands manageable, which explains why the cumulative error remains vastly lower than a single, sweeping volumetric jump.Solvent matrix suppression phenomena
Do not assume pure water behaves like complex environmental samples. When you learn how to dilute 1000 ppm to 10 ppm in a matrix like saltwater or industrial wastewater, background interference changes everything.
The viscosity of the solvent alters how liquid drains from glass walls. Highly viscous matrices require longer drainage times, meaning a rapid dispensation leaves valuable milligrams behind. Always match the matrix of your calibration standards to the exact profile of your experimental blanks to ensure uniform behavior.
Frequently asked questions
Can I use tap water when figuring out how to dilute 1000 ppm to 10 ppm?
Absolutely not, unless you want to sabotage your analytical baseline before the experiment even starts. Municipal tap water contains background minerals like calcium, magnesium, and chlorine, often fluctuating between 150 ppm and 300 ppm of total dissolved solids. Introducing these unknown ionic variables completely invalidates your mathematical ratios. You must utilize Deionized Water (DI) with 18.2 megohm-cm resistivity to ensure your solvent contributes zero trace elements to the final mixture.How long will a 10 ppm diluted solution remain stable?
Low concentration standards degrade rapidly due to a phenomenon known as container wall adsorption. At a trace level of 10 ppm, individual solute ions actively bind to the microscopic active sites on the interior surface of glass or plastic storage bottles. A solution left on the shelf for more than 48 hours will often measure significantly lower than its original formulation. For maximum reproducibility, prepare your final working concentrations fresh on the day of analysis and store them in inert fluorinated ethylene propylene containers.What is the maximum allowable error when measuring the initial 1000 ppm stock?
Your final accuracy is rigidly bound to the tolerances of your starting equipment. Utilizing a standard 10-milliliter Class A volumetric pipette introduces a strict manufacturing tolerance of plus or minus 0.02 milliliters. If you deviate beyond this envelope during your initial draw, the geometric progression of a serial dilution will carry that deviation through every subsequent phase. Treat the primary measurement as the most vulnerable link in your analytical chain.Definitive synthesis on precision metrics
Relying blindly on automated pipettes without verifying their calibration is a recipe for scientific fiction. We must stop treating trace-level preparation as a casual kitchen recipe. The math behind understanding how to dilute 1000 ppm to 10 ppm is simple arithmetic, yet the physical execution requires flawless laboratory discipline. If your technique is sloppy, your data is garbage. (And let us be honest, automated tools often mask poor manual technique.) Invest the necessary time into executing a two-step serial progression rather than taking shortcut direct leaps. True accuracy is won or lost in the meticulous alignment of the meniscus line.